LJ Fluid#
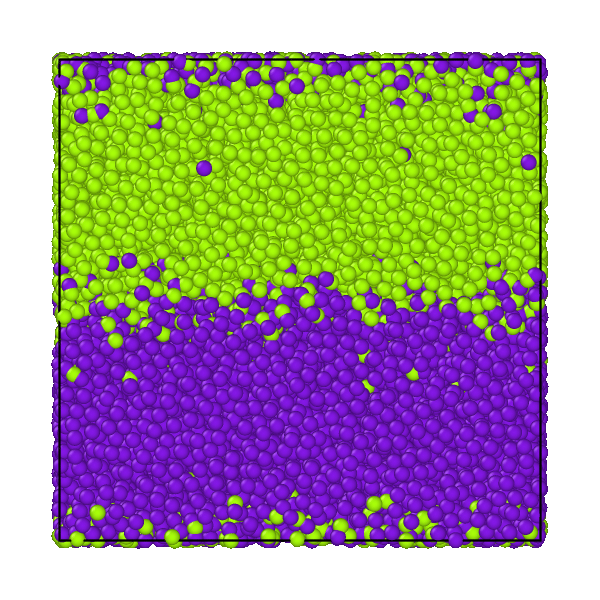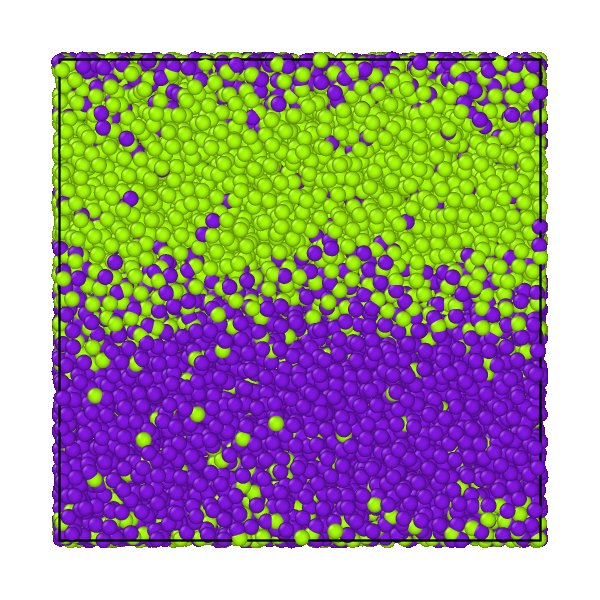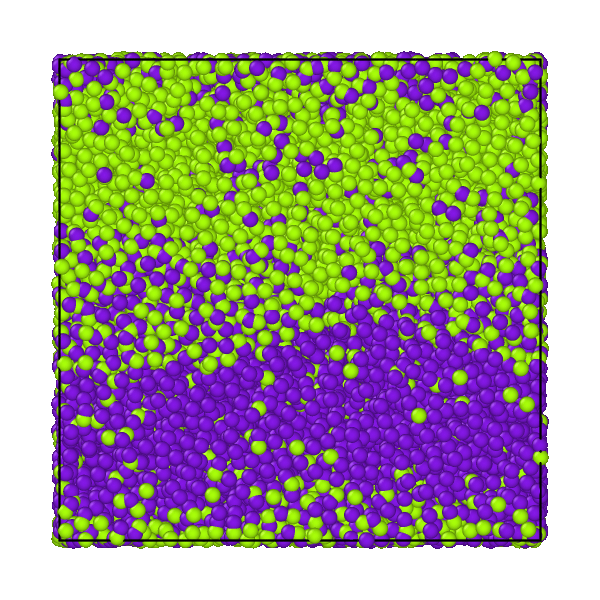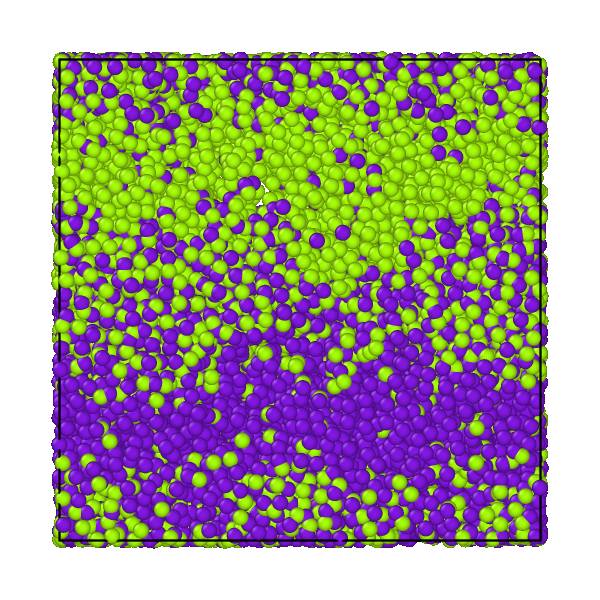
In this hands-on guide, we’ll use LAMMPS to simulate a mixture of two fluids (binary fluid) and study the phase separation. The fluids are modeled as Lennard-Jones particles confined within a cubic box with continuous boundaries (periodic boundary conditions). We’ll employ a Langevin thermostat to control the temperature, and calculate fundamental properties like potential and kinetic energies. This tutorial serves as a stepping stone, showcasing essential elements of molecular dynamics simulations, including setting up the system, minimizing energy, solving the equations of motion, and visualizing the resulting particle trajectories.
Introduction to simple LAMMPS script#
In order to go through step by step lets consider the simulation framework. Simulation might contains several files. But here we mainly focus on:
a)
run.in– it cpntains mostly all the informationsb)
system.data– particles, positions, velocities, bonds, angles, information
1) Initialization: units, dimension, atom_style, pair_style boundary etc.
2a) System definition: either load
system.dataorcreate_atoms2b) Bond and Angle information:
pair_coeff,angle_coeff,mass3) Minimization: to avoid initial overlaps
4) Visualization: Check whether particles are in the box etc.
5) Dummy Run: Initially run for small timesteps i.e., \(\sim 10^5\)
6) Reset time:
write_data,reset_timesteps7) Final Run:
With the help of ChatGPT you can built your first simulation
`Promt:`
I am a beginner looking to learn the LAMMPS package from scratch. Could you provide me with step-by-step instructions on how to set up and run basic simulations? Specifically, I'd like:

- 1. Detailed instructions on installing LAMMPS on my operating system.
- 2. An overview of the structure of a LAMMPS input script.
- 3. A couple of simple example simulations, including the full input script for each.
- 4. Instructions on how to run these simulations.
- 5. Tips on how to analyze the output and visualize the results.
- 6. Recommendations for further learning resources or tutorials.

Please provide the input scripts and any necessary commands to run the simulations.
system.data: an example of lammps generated system file
LAMMPS data file via write_data, version 3 Mar 2020, timestep = 0

200 atoms
5 atom types
199 bonds
1 bond types
198 angles
1 angle types

0.0000000000000000e+00 5.0000000000000000e+01 xlo xhi
0.0000000000000000e+00 5.0000000000000000e+01 ylo yhi
0.0000000000000000e+00 5.0000000000000000e+01 zlo zhi

Masses

1 1
2 1
3 1
4 1
5 1

Atoms # full

35 1 4 0.0000000000000000e+00 4.6927442356735957e-04 2.5917304644577136e+01 9.2446343214115345e-01 1 0 1
24 1 3 0.0000000000000000e+00 4.4738596798170382e+00 2.7817888523819551e+01 5.6717976560832186e+00 1 0 1
200 1 1 0.0000000000000000e+00 5.9147516604228179e-01 1.8162531643029414e+01 4.7691758446364831e+00 1 0 1
166 1 5 0.0000000000000000e+00 1.4206564943257913e-01 2.5387383075812657e+01 4.9959602477906309e+01 1 0 0
168 1 3 0.0000000000000000e+00 5.9831302455988533e-01 2.7059222183708062e+01 4.9375698118072819e+01 1 0 0
167
.
.
.
198 1 3 0.0000000000000000e+00 4.9886372673067399e+01 1.9279039503505089e+01 3.3634226785821535e+00 0 0 1
188 1 3 0.0000000000000000e+00 4.9545910789376187e+01 2.5180567455466711e+01 4.8987638671791863e+01 0 0 0
89 1 2 0.0000000000000000e+00 4.9901224571757616e+01 1.5141601528783990e+01 4.6754960184097342e+01 0 0 0
114 1 3 0.0000000000000000e+00 4.9958704483647480e+01 2.1674563605430009e+01 1.3738041526491067e+01 0 0 1

Velocities

35 -8.7770147801311138e-01 1.5101693717367606e+00 6.0446096933556570e-01
24 -9.9424974102245878e-01 1.3375187903264580e+00 2.5207278347123672e+00
200 1.1303805073183029e+00 6.5838908974304966e-01 1.6455895349853347e+00
166 -2.1711562280283929e+00 5.5739576720902884e-01 -2.4101260924931348e-01
168 3.9698835008419225e-02 5.8955871401596471e-01 2.3171435957398314e+00
.
.
.
188 -1.8696257894080665e-01 -1.1207437904225968e+00 -3.9930018404493181e-03
89 2.9740209786705213e-01 7.0494417493173855e-01 7.4736174728733051e-01
114 -4.7738544094380891e-01 5.8646358115481168e-01 -1.3337631131086364e-01


Bonds

1 1 35 36
2 1 24 25
3 1 166 167
4 1 168 169
5 1 167 168
.
.
198 1 89 90
199 1 114 115

Angles

1 1 34 35 36
2 1 23 24 25
3 1 165 166 167
4 1 167 168 169
.
.
What do these numbers mean?#
system.data: an example of lammps generated system file
LAMMPS data file via write_data, version 3 Mar 2020, timestep = 0

200 atoms
5 atom types
199 bonds
1 bond types
198 angles
1 angle types

X_low X_high xlo xhi :box dimension along x
Y_low Y_high ylo yhi :box dimension along y
Z_low Z_high zlo zhi :box dimension along z

Masses

atom_type mass
1 1
2 1

Atoms # full

atom_ID molecular_ID atom_type charge x y z ix iy iz(image in the periodic boundary)
1 1 1 0 2 3 4 0 0 0
2 1 2 0 3 5 1 0 0 0

Velocities

atom_ID vx vy vz
1 0.5 0.4 0.33

Bonds

ID Bond_type atom1 atom2 (atom1 atom2 are sequentially connected via a bond)
1 1 1 2
2 1 2 3

Angles

ID Angle_type atom1 atom2 atom3 (atom1 atom2 and atom3 are sequentially connected)
1 1 1 2 3
2 1 2 3 4
Important
Please make sure you always keep the format fixed No space between different segements might leads to syntax error
Lets pick an example of LJ fluid.
What are the parameters we need to specify
a) Numer of particles
b) Non-bonded interactions
c) Mass
d) Bonds, angle, dihedrals etc.
e) What type of bonds, angles, dihedrals etc. and their corresponding parameters
f) Boundaries
g) How long simulation would run
h) timesteps, otherwise defult
i) How frequently dump the configurations
Method 1: run.in: LAMMPS script for a 10-particle LJ fluid
units lj
dimension 3
atom_style full
pair_style lj/cut 1.112
boundary p p p

region          simulation_box block 0 20 0 20 0 20
create_box      1 simulation_box
create_atoms    1 random 10 12345 NULL overlap 2.0 maxtry 50

mass            1 1
pair_coeff      * * 1.0 1.0


thermo          100
thermo_style    custom step temp pe ke etotal press
timestep        0.005
fix             mynve all nve
fix             mylgv all langevin 1.0 1.0 0.1 1530917
run             10000
reset_timestep 0
dump            mydmp all atom 1000 dump.lammpstrj
run             1000000
log file
.
.
.
.
    999100   1.3074207      0              1.765018       1.765018       0.0014708483
    999200   1.4026846      0              1.8936242      1.8936242      0.0015780201
    999300   0.72046866     0              0.9726327      0.9726327      0.00081052725
    999400   0.78655881     0              1.0618544      1.0618544      0.00088487866
    999500   1.0822006      0              1.4609708      1.4609708      0.0012174757
    999600   0.90615892     0              1.2233145      1.2233145      0.0010194288
    999700   0.985891       0              1.3309528      1.3309528      0.0011091274
    999800   1.2901643      0              1.7417218      1.7417218      0.0014514348
    999900   0.67689256     0              0.91380495     0.91380495     0.00076150413
   1000000   0.78634406     0              1.0615645      1.0615645      0.00088463707
Loop time of 19.87 on 8 procs for 1000000 steps with 10 atoms

Performance: 21741277.904 tau/day, 50327.032 timesteps/s, 503.270 katom-step/s
80.1% CPU use with 8 MPI tasks x no OpenMP threads

MPI task timing breakdown:
Section |  min time  |  avg time  |  max time  |%varavg| %total
---------------------------------------------------------------
Pair    | 0.054741   | 0.057883   | 0.064233   |   1.2 |  0.29
Bond    | 0.04021    | 0.042227   | 0.044459   |   0.8 |  0.21
Neigh   | 0.34606    | 0.35955    | 0.37062    |   1.3 |  1.81
Comm    | 7.498      | 8.0589     | 8.8711     |  16.4 | 40.56
Output  | 0.57603    | 0.61785    | 0.73808    |   6.0 |  3.11
Modify  | 0.16301    | 0.17659    | 0.1904     |   2.4 |  0.89
Other   |            | 10.56      |            |       | 53.13

Nlocal:           1.25 ave           4 max           0 min
Histogram: 4 0 1 0 0 1 0 1 0 1
Nghost:          2.125 ave           4 max           0 min
Histogram: 2 0 0 0 0 2 0 3 0 1
Neighs:              0 ave           0 max           0 min
Histogram: 8 0 0 0 0 0 0 0 0 0

Total # of neighbors = 0
Ave neighs/atom = 0
Ave special neighs/atom = 0
Neighbor list builds = 78413
Dangerous builds = 0
Total wall time: 0:00:20
Method 2: run.in: LAMMPS script read_data from system_specific.data file
units lj
dimension 3
atom_style full
pair_style lj/cut 1.112
boundary p p p

read_data        system_box.data

pair_coeff      * * 1.0 1.0


thermo          100
thermo_style    custom step temp pe ke etotal press
timestep        0.005
fix             mynve all nve
fix             mylgv all langevin 1.0 1.0 0.1 1530917
run             10000
reset_timestep 0
thermo          1000
dump            mydmp all atom 1000 dump.lammpstrj
run             1000000
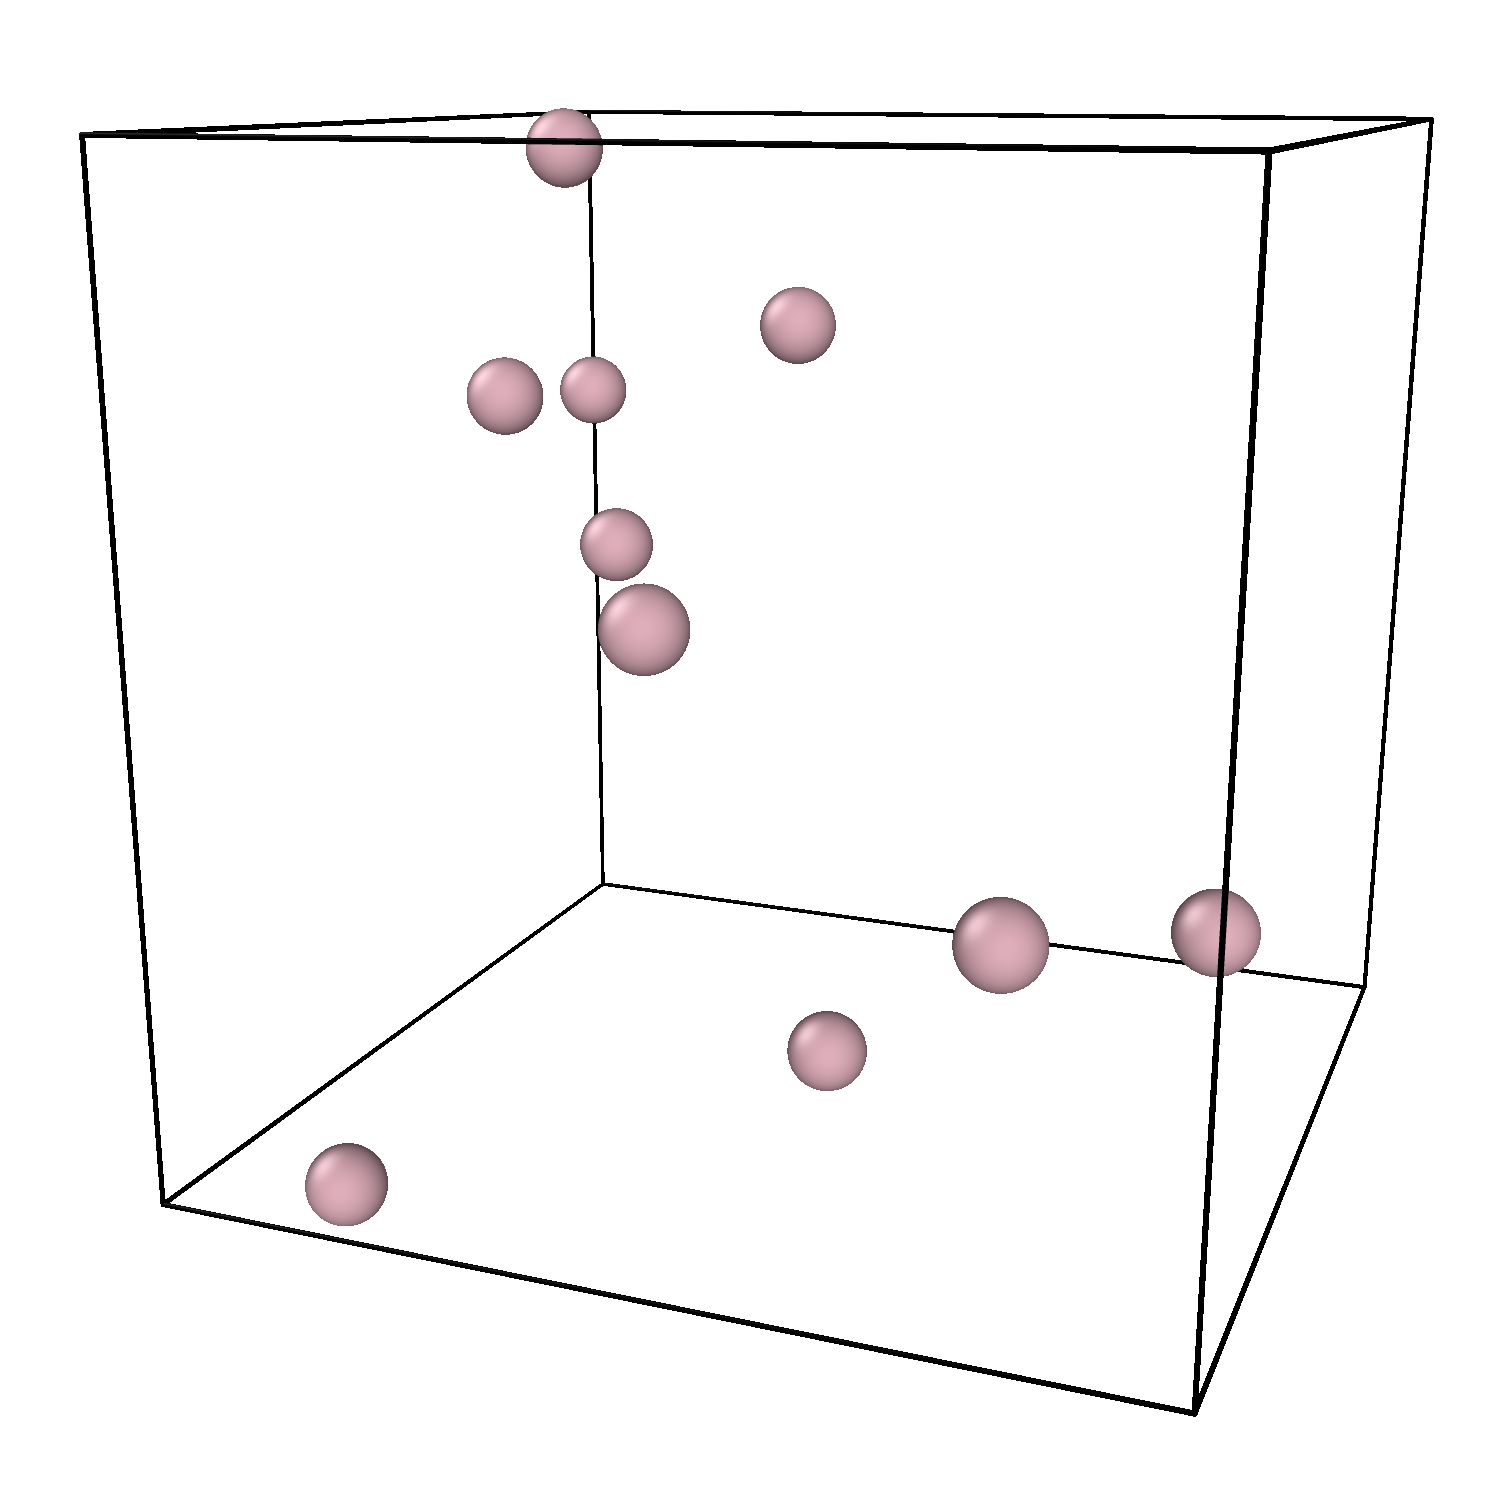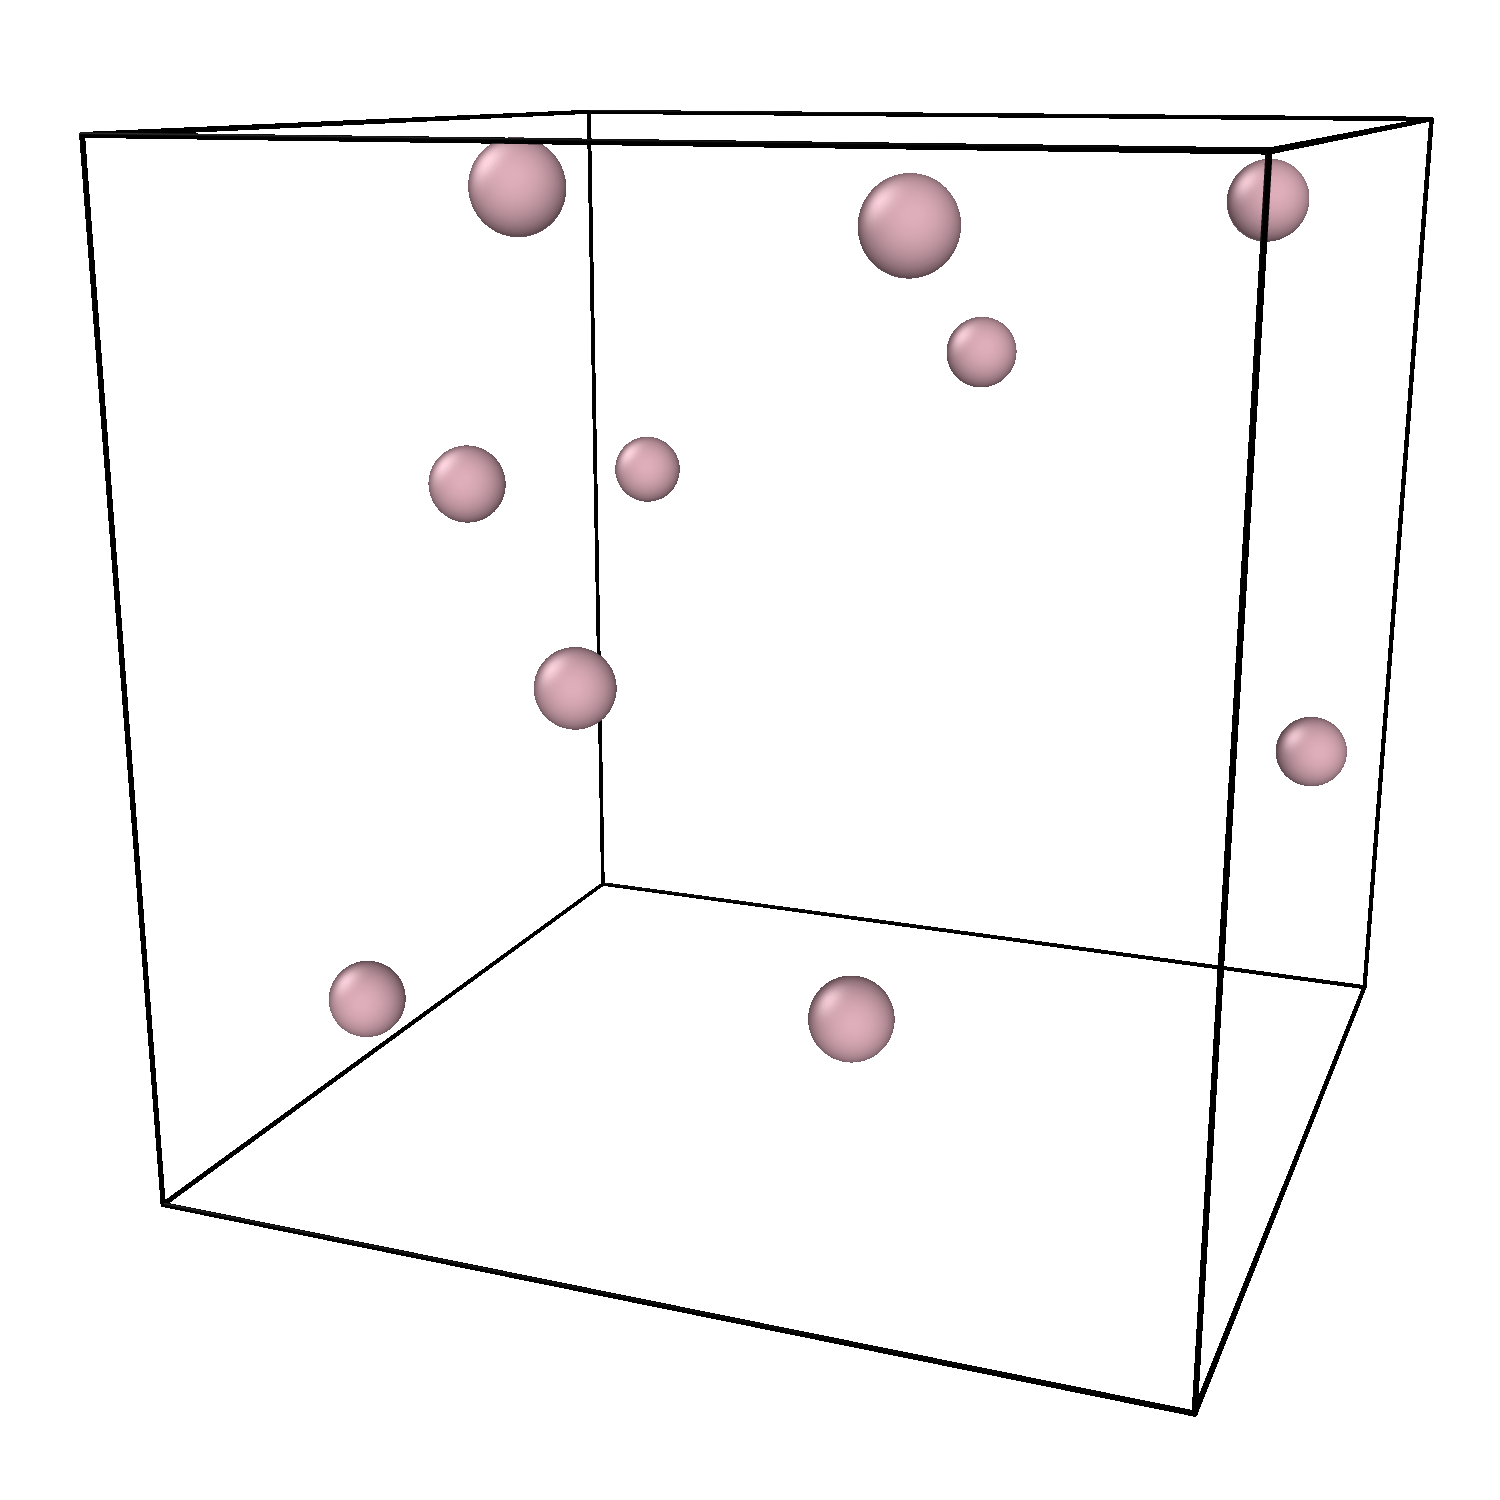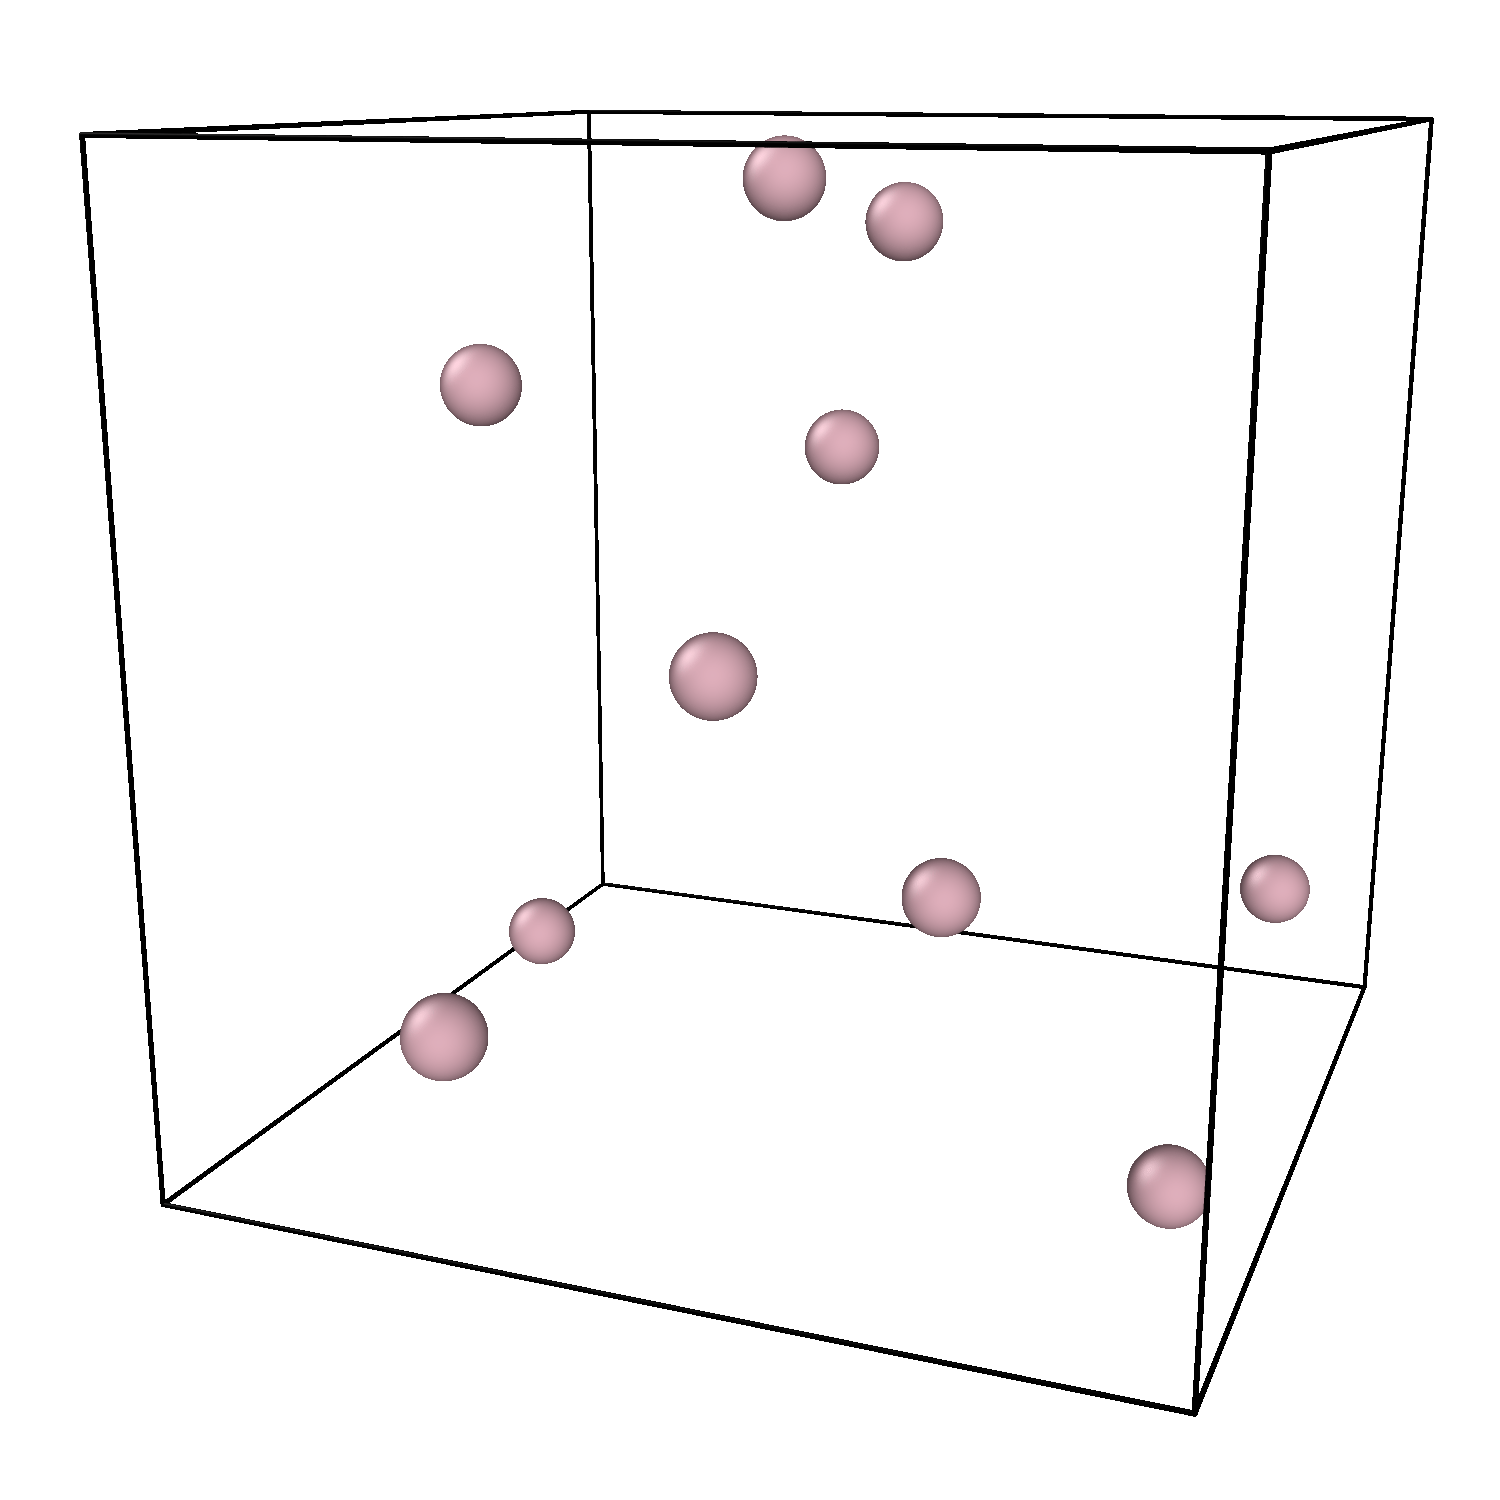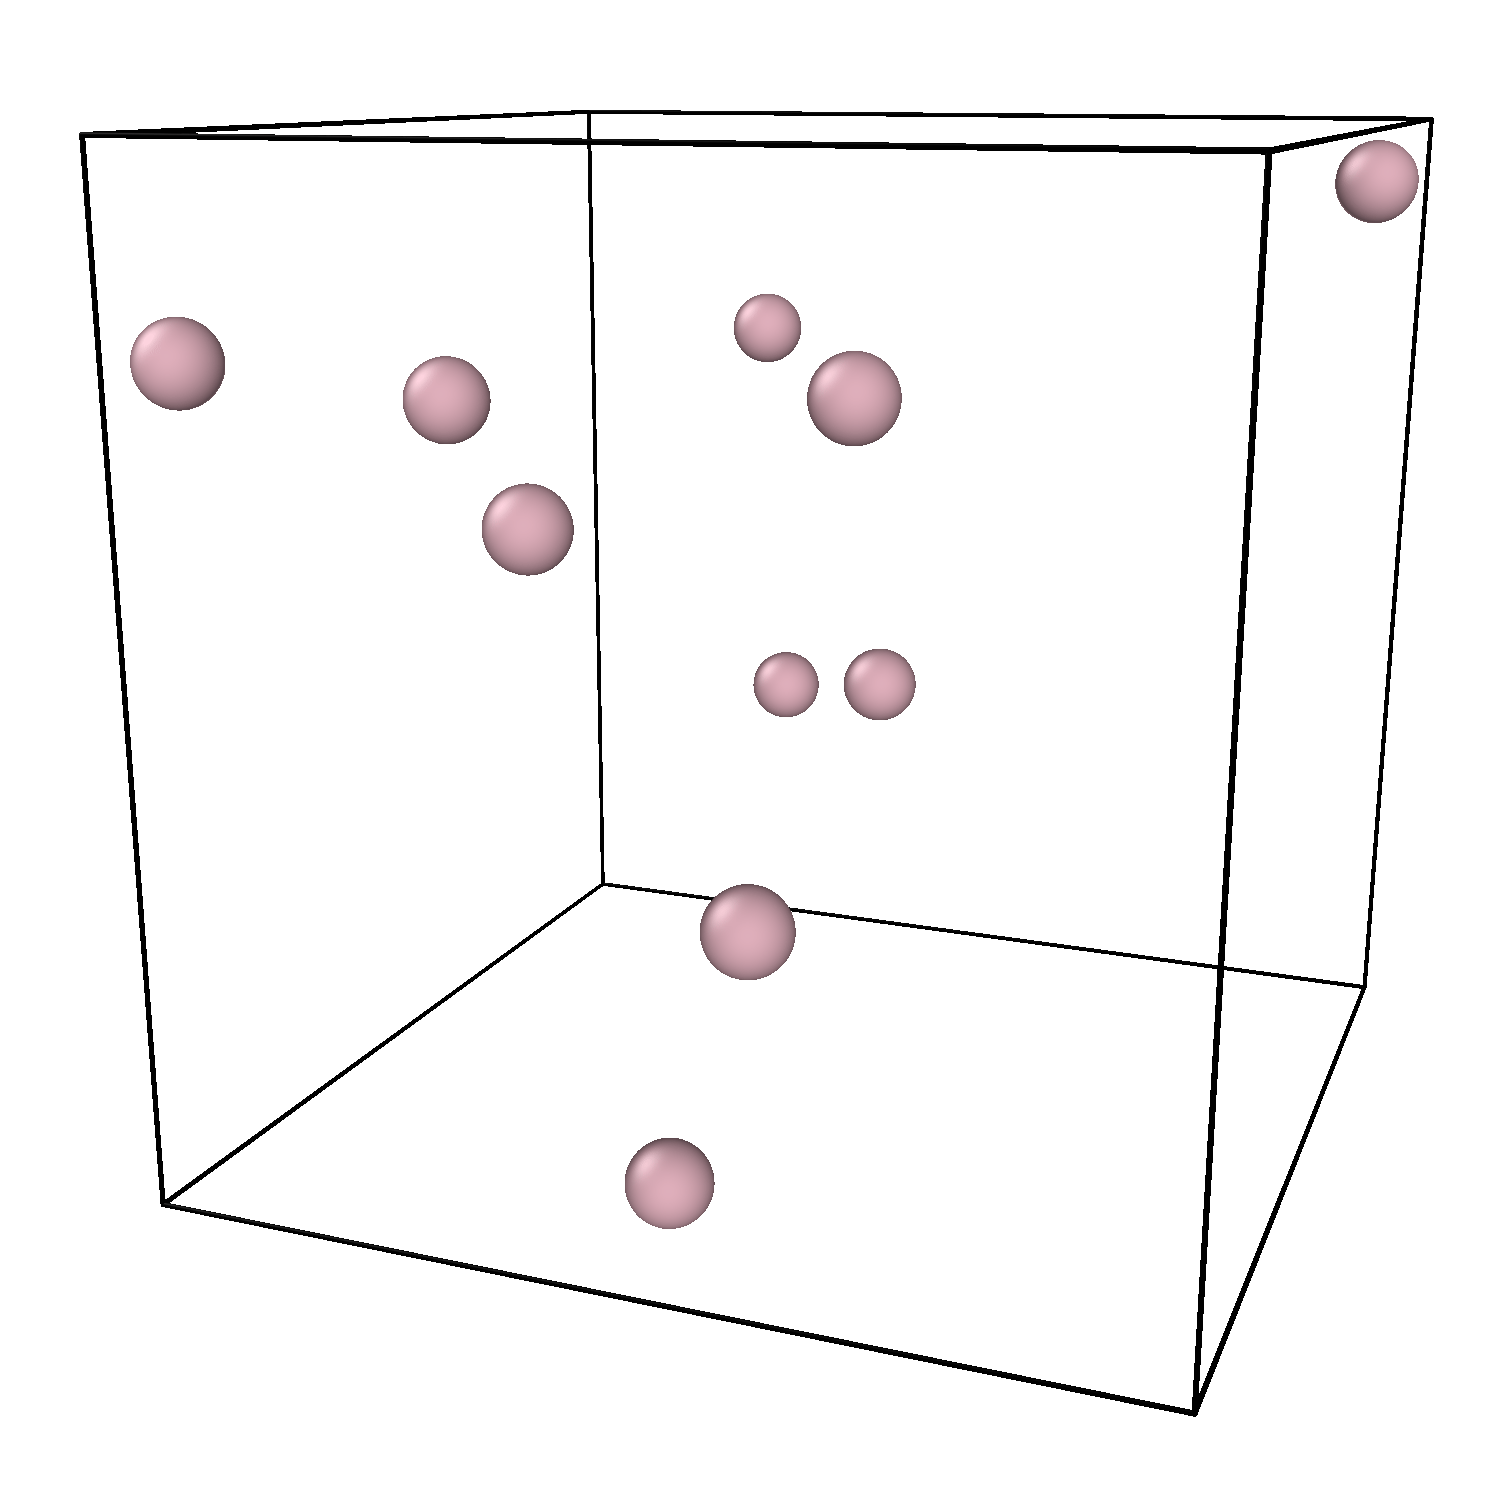
Method 2: system_box.data: LAMMPS script system_box.data file
LAMMPS data file via write_data, version 3 Mar 2020

10 atoms
1 atom types

0 10 xlo xhi
0 10 ylo yhi
0 10 zlo zhi

Masses

1 1

Atoms # full

1 1 1 0 1 1 1
2 1 1 0 2 2 2
3 1 1 0 3 1 1
4 1 1 0 4 2 2
5 1 1 0 5 1 1
6 1 1 0 6 2 2
7 1 1 0 7 1 1
8 1 1 0 8 2 2
9 1 1 0 9 1 1
10 1 1 0 1 5 2
Method 3: run.in: LAMMPS script read_data from system_specific.data file populate the box with molecules replication
units           lj
dimension       3
atom_style      full
pair_style      lj/cut 1.112
boundary        p p p

read_data       system_box.data
replicate       3 3 3

pair_coeff      * * 1.0 1.0


thermo          100
thermo_style    custom step temp pe ke etotal press
timestep        0.005
fix             mynve all nve
fix             mylgv all langevin 1.0 1.0 0.1 1530917
run             10000
reset_timestep  0
thermo          1000
dump            mydmp all atom 1000 dump.lammpstrj
run             1000000
Method 3: system_box.data: LAMMPS script system_box.data file specify only one molecule and replicate it
LAMMPS data file via write_data, version 3 Mar 2020,

1 atoms
1 atom types

0 2 xlo xhi
0 2 ylo yhi
0 2 zlo zhi

Masses

1 1

Atoms # full

1 1 1 0 1 1 1
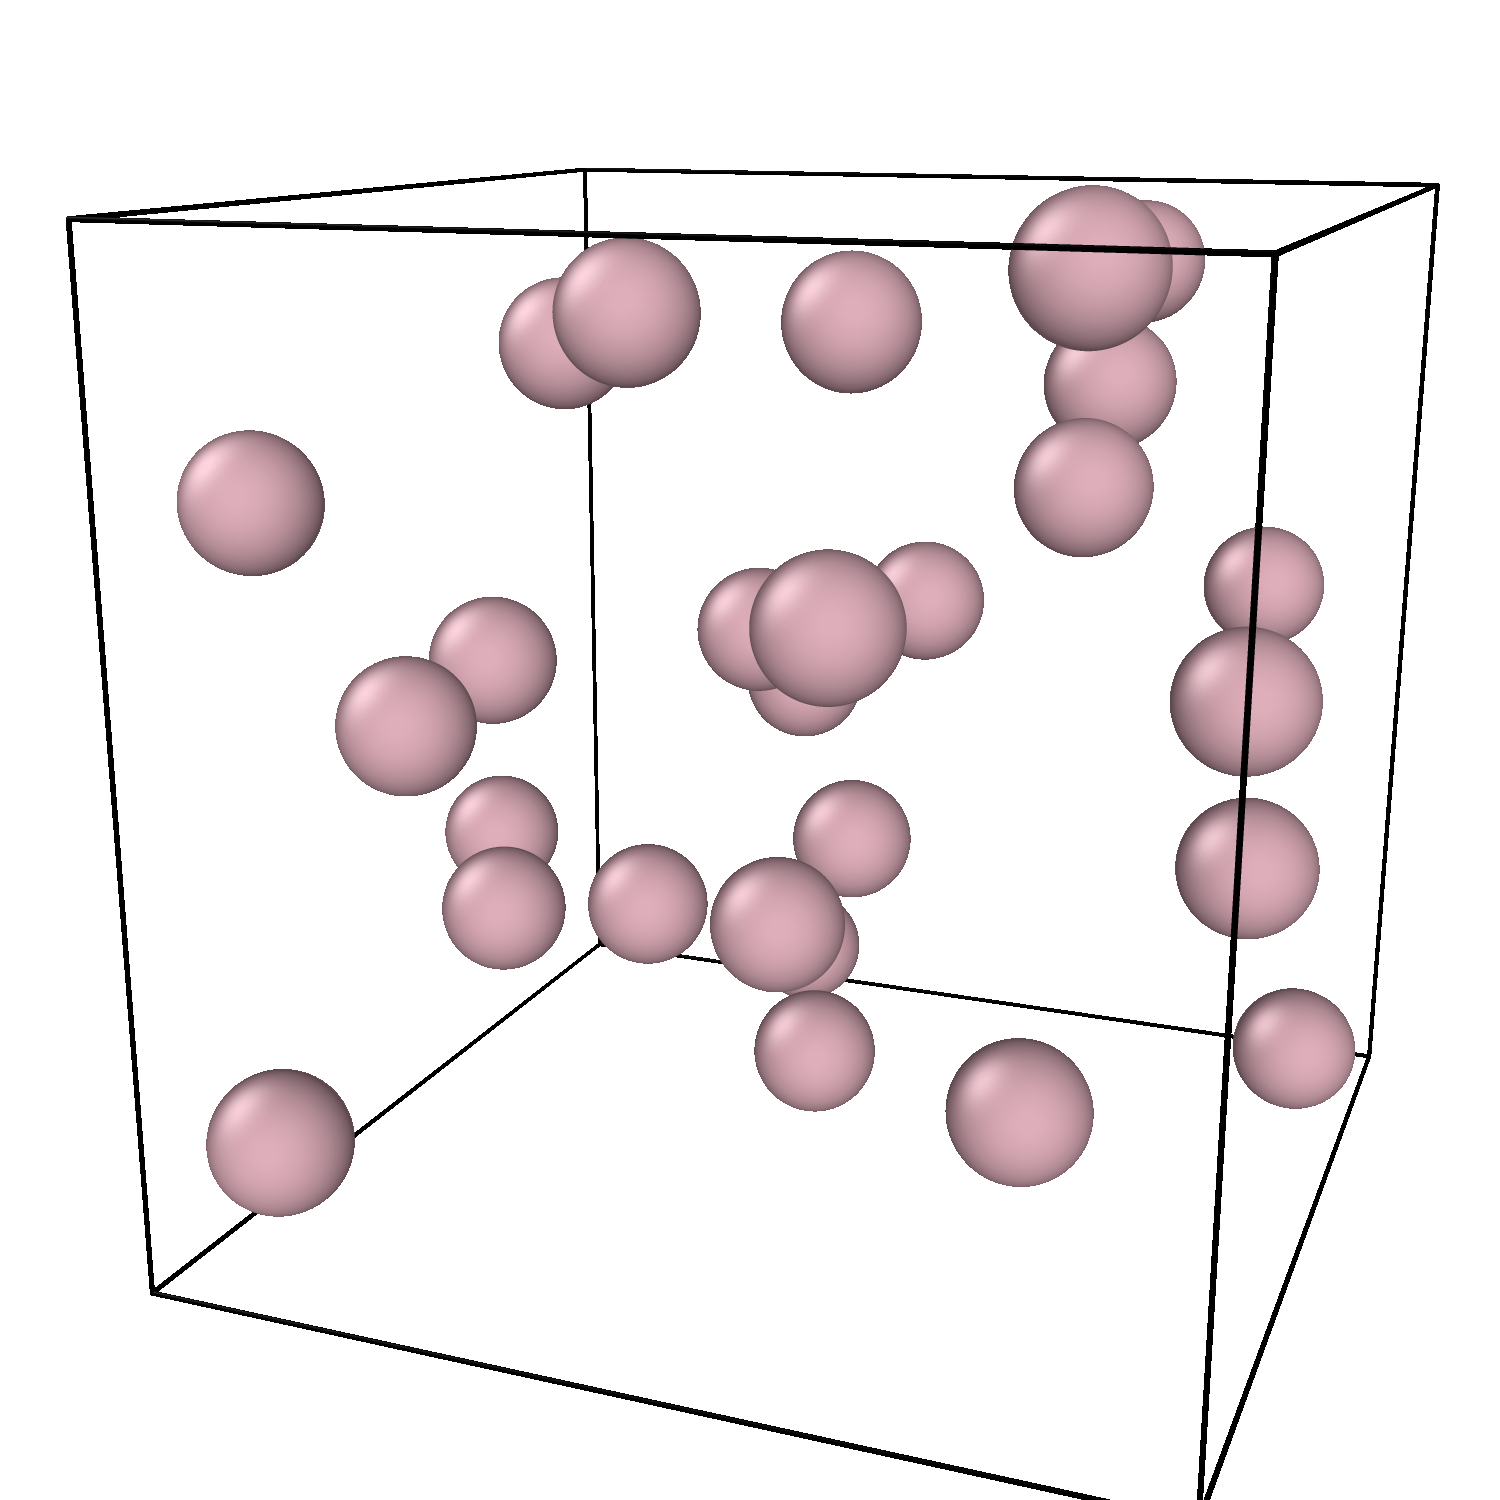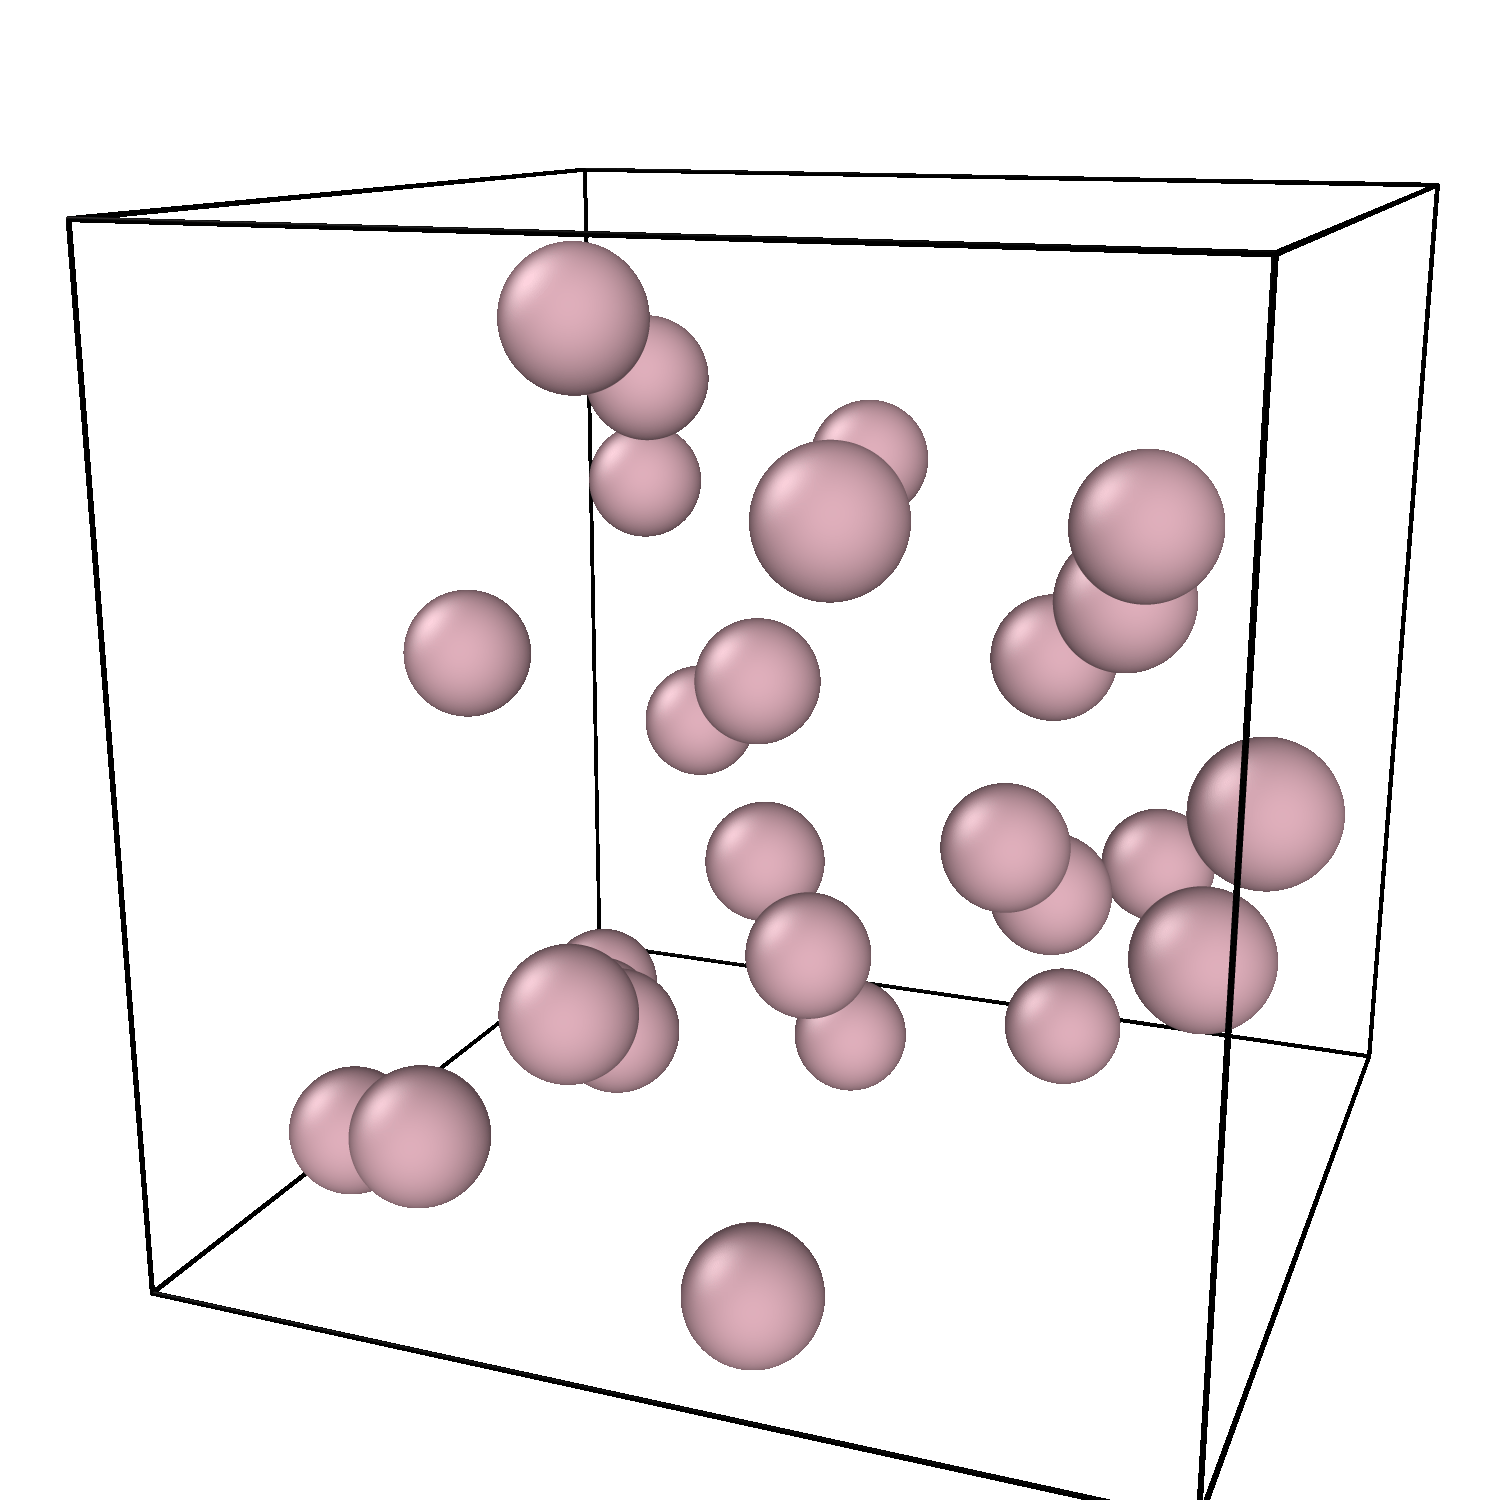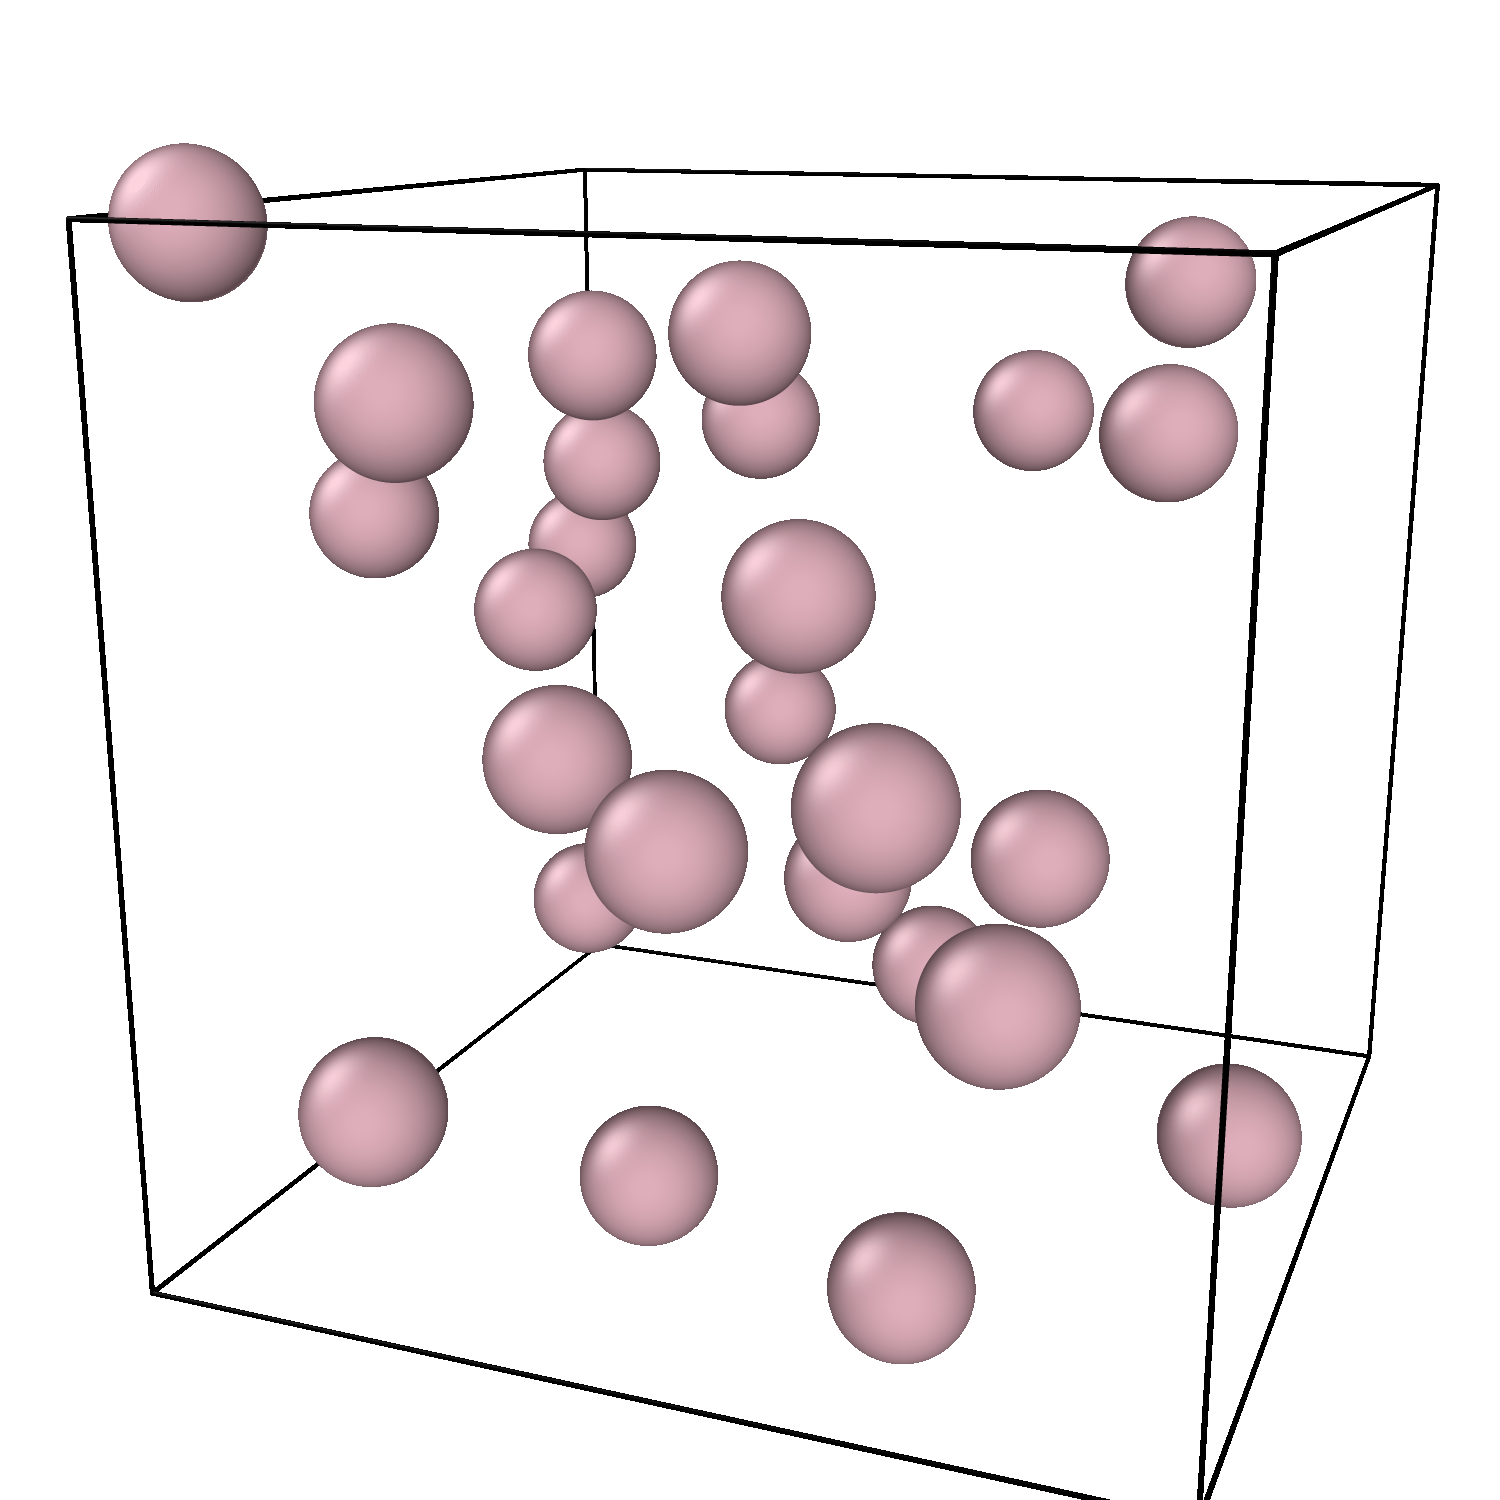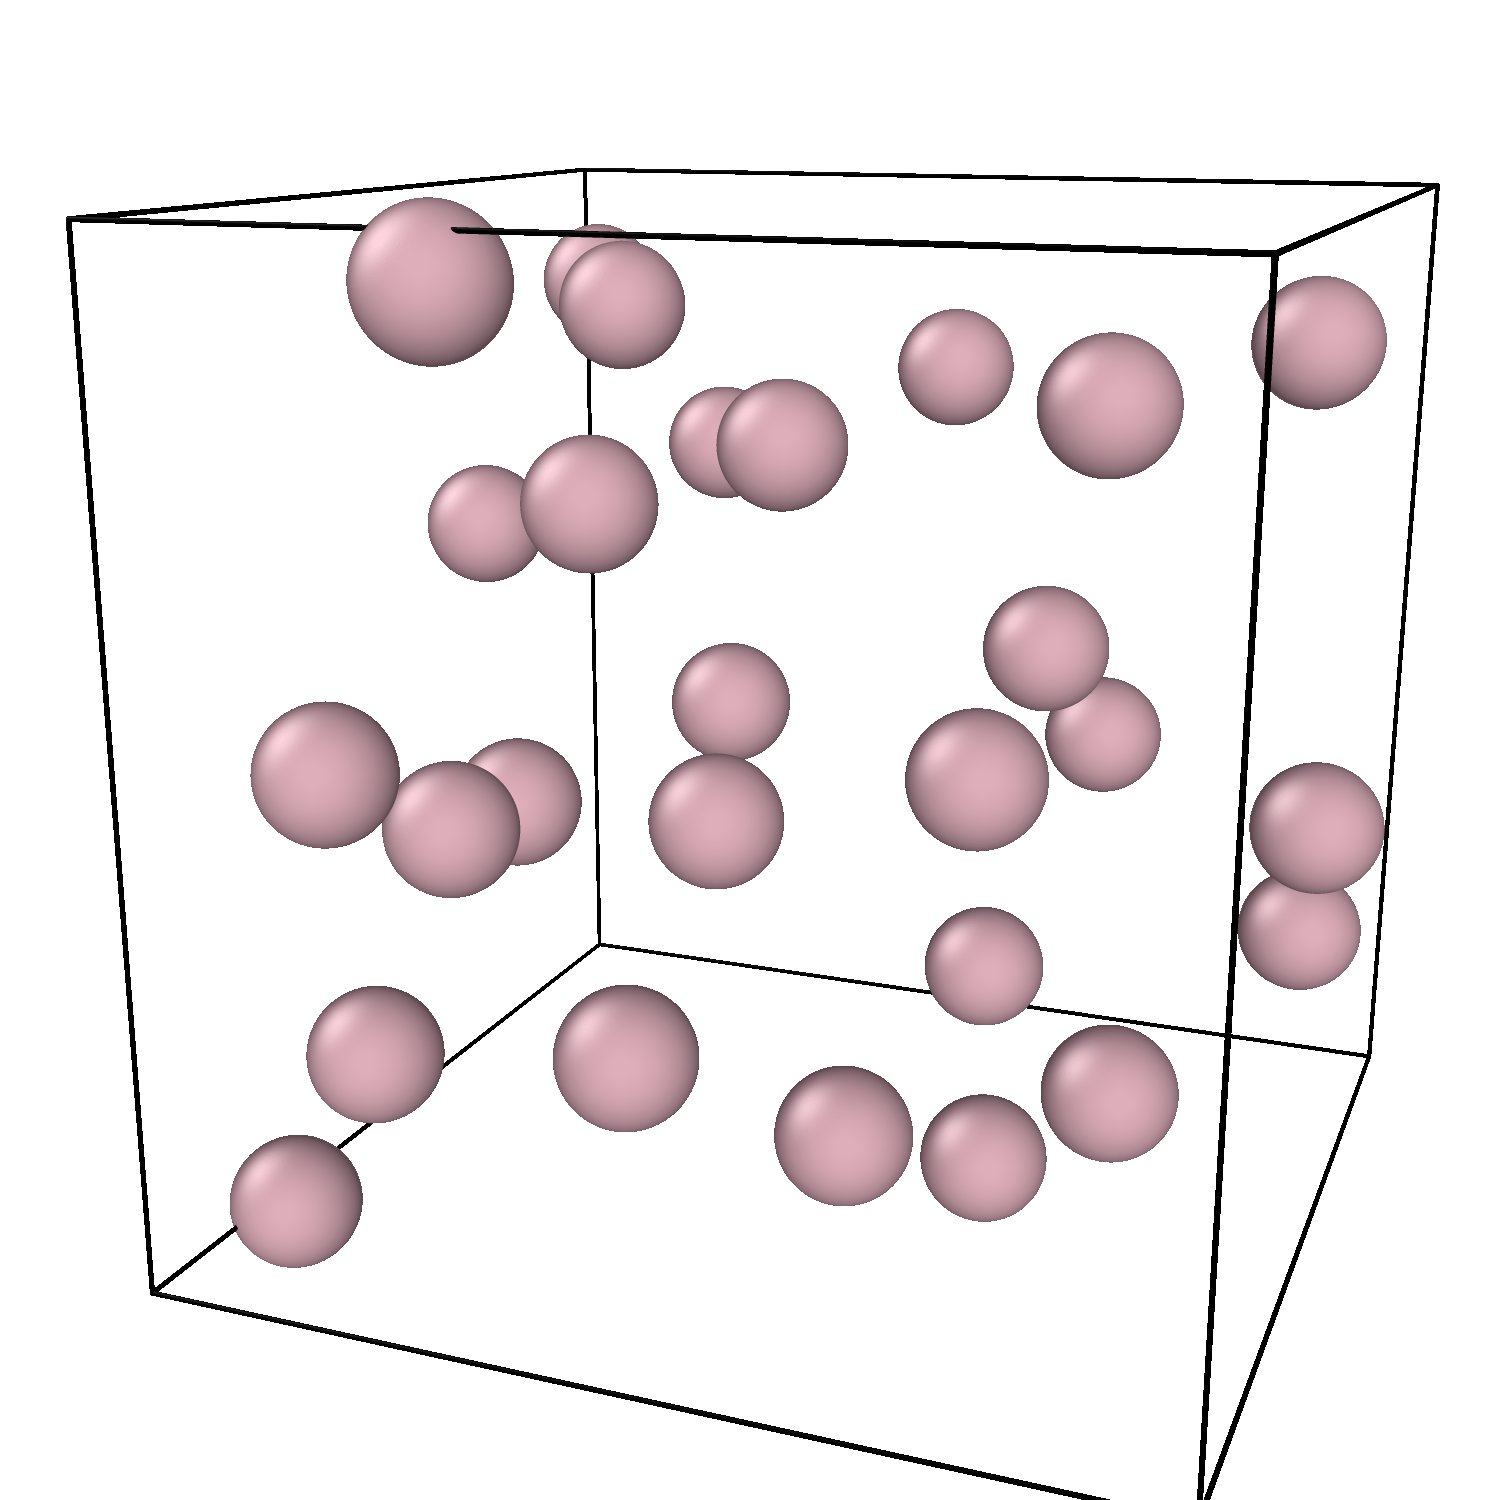
Lets pick an example of Binary mixture of LJ fluid.
Binary immiscible fluid
a) Intermolecular interaction between Phase A and Phase A attract each other
b) Intermolecular interaction between Phase B and Phase B attract each other
c) Phase A and Phase B dislike each other
d) Box is filled with Phase A and Phase B around 50%:50%
run.in: LAMMPS script for a binary mixture of LJ fluid
units           lj
dimension       3
atom_style      full
pair_style      lj/smooth/linear 3.5
boundary        p p p

region          simulation_box block 0 30 0 30 0 30
region          simulation_box_sidea block 0 30 0 30 0 14.5
region          simulation_box_sideb block 0 30 0 30 15 30
create_box      2 simulation_box

lattice         fcc 1.0
create_atoms    1 region simulation_box_sidea ratio 0.63 5478952
create_atoms    2 region simulation_box_sideb ratio 0.63 5478952

mass            * 1
pair_coeff      1 1 1.0 1.0  #AA
pair_coeff      1 2 1.0 0.5  #AB
pair_coeff      2 2 1.0 1.0  #BB
write_data 		  system_formation.data

thermo          500
thermo_style    custom step temp pe ke etotal density press
timestep        0.005
fix             mynve all nve
fix             mylgv all langevin 1.0 1.0 0.1 1530917
run             10000
write_data 		  system_after_equilibrium.data
reset_timestep  0
thermo          5000
dump            mydmp all atom 1000 dump.lammpstrj
run             1000000

write_data      system_after_longtime.data